library(sf)
library(dplyr)
library(ggplot2)
library(patchwork)
library(spatialdataR)
library(SpatialData.data)
library(SpatialData.plot)MCMICRO
spatial proteomics
MCMICRO
In this demo, we’ll analyze 12-plex CyCIF data on a human lung adenocarcinoma core, as output by MCMICRO (Schapiro et al. 2022), which stems from a larger tissue microarray (TMA); included are also cell segmentation outputs from both UnMICST and Ilastik, respectively, as well as corresponding channel x cell protein expression matrices (here, corresponding to mean pixel intensities).
preamble
(sd <- SD.data_load("LungAdenocarcinomaMCMICRO"))class: SpatialData
- images(1): exemplar-001_image
- labels(2): exemplar-001_cell exemplar-001_nuclei
- tables(2):
- exemplar-001--ilastik_cell (12,11607) [exemplar-001_cell]
- exemplar-001--unmicst_cell (12,11170) [exemplar-001_cell]
coordinate systems(1):
- global(3): exemplar-001_image exemplar-001_cell exemplar-001_nucleiLet’s start by viewing available channels():
cat(channels(image(sd)))DNA_6 ELANE CD57 CD45 DNA_7 CD11B SMA CD16 DNA_8 ECAD FOXP3 NCAMFor consistency, we will fix a color to use for each channel:
pal <- c(
    DNA_6="grey",
    DNA_7="grey",
    DNA_8="grey",
    CD57="maroon",  # NK
    FOXP3="yellow", # Treg
    SMA="red",      # stromal
    NCAM="orange",  # myocyte
    CD11B="blue",   # dendritic
    CD16="cyan",    # neutrophil
    ELANE="purple", # neutrophil
    ECAD="green",   # epithelial
    CD45="magenta") # lymphocyte
The original image includes a superfluous empty region, which we’ll remove:
sd <- crop(sd, list(xmin=0, xmax=2500, ymin=600, ymax=3200))Now, let’s generate a composite image of few selected channels:
ch <- c("CD16", "CD45", "FOXP3", "ELANE", "ECAD", "SMA")
plotSpatialData(sd) + plotImage(ch=ch, c=pal[ch])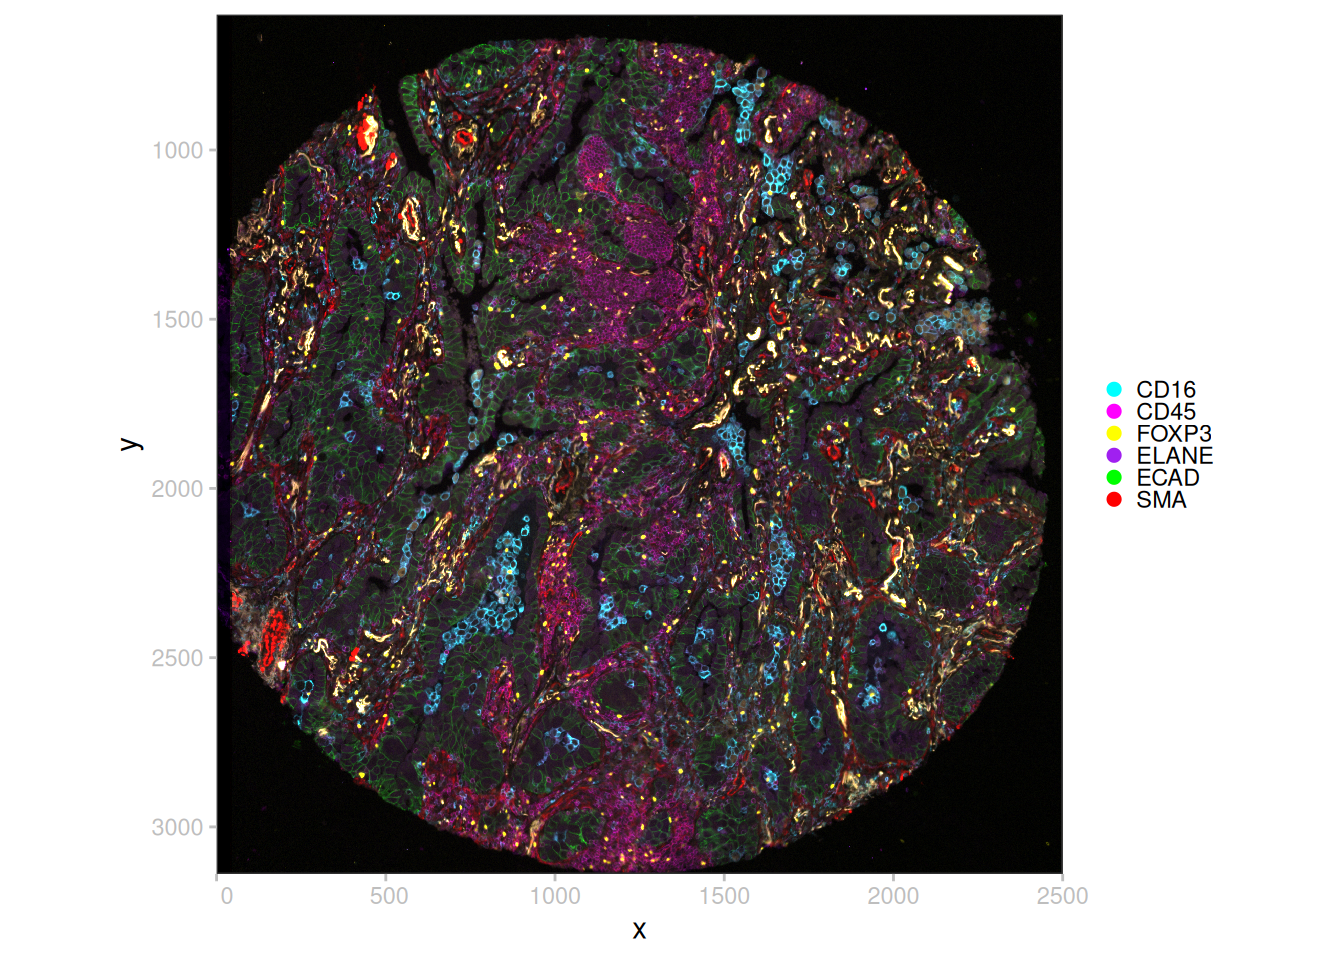
We can also visualize channels individually (here, using a smaller region):
bb <- list(
    xmin=1600, xmax=2400,
    ymin=1200, ymax=2000)
sp <- crop(sd, bb)

ch <- channels(image(sp))
ps <- lapply(ch, \(.) plotSpatialData(sp) + plotImage(ch=., c=pal[.]) + ggtitle(.))

wrap_plots(ps, nrow=3) &
    theme_void() & theme(
        legend.position="none",
        plot.title=element_text(hjust=0.5))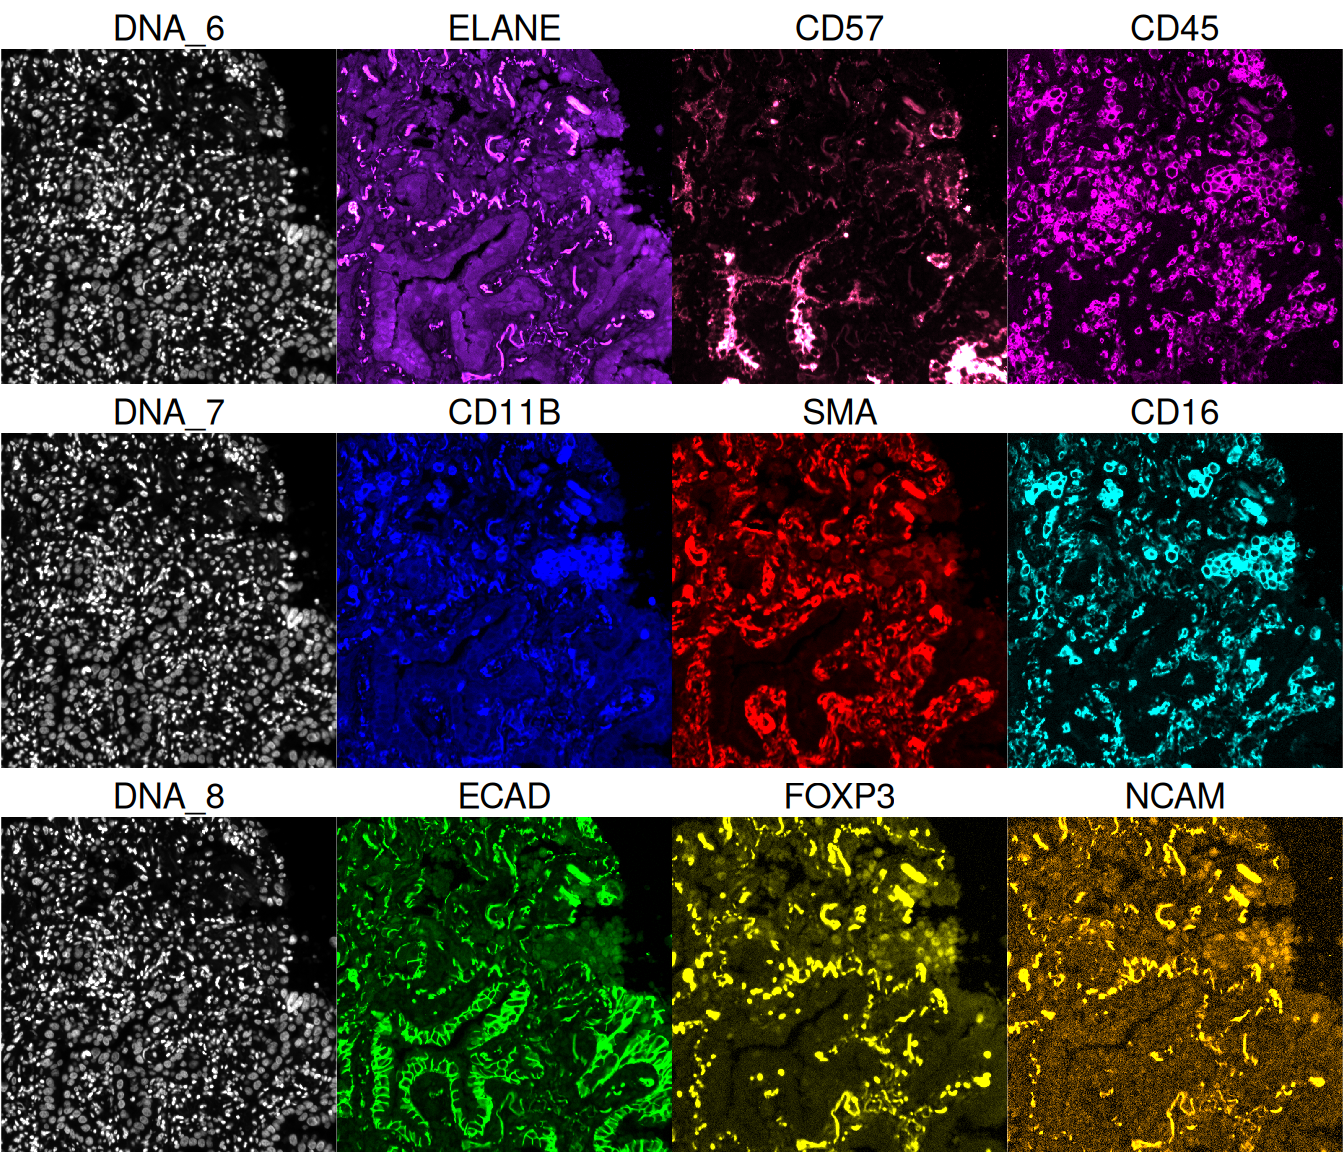
(se <- table(sd))class: SingleCellExperiment
dim: 12 11607
metadata(0):
assays(1): X
rownames(12): DNA_6 ELANE ... FOXP3 NCAM
rowData names(9): channel_number cycle_number ... exposure remove
colnames(11607): 0 1 ... 11605 11606
colData names(7): Area MajorAxisLength ... Extent Orientation
reducedDimNames(1): spatial
mainExpName: NULL
altExpNames(0):appendix
Notesession
sessionInfo()R version 4.6.2 (2026-10-29)
Platform: x86_64-pc-linux-gnu
Running under: Ubuntu 26.04.1 LTS
Matrix products: default
BLAS: /usr/lib/x86_64-linux-gnu/openblas-pthread/libblas.so.3
LAPACK: /usr/lib/x86_64-linux-gnu/openblas-pthread/libopenblasp-r0.3.32.so; LAPACK version 3.12.0
locale:
[1] LC_CTYPE=C.UTF-8 LC_NUMERIC=C LC_TIME=C.UTF-8
[4] LC_COLLATE=C.UTF-8 LC_MONETARY=C.UTF-8 LC_MESSAGES=C.UTF-8
[7] LC_PAPER=C.UTF-8 LC_NAME=C LC_ADDRESS=C
[10] LC_TELEPHONE=C LC_MEASUREMENT=C.UTF-8 LC_IDENTIFICATION=C
time zone: UTC
tzcode source: system (glibc)
attached base packages:
[1] stats graphics grDevices datasets utils methods base
other attached packages:
[1] SpatialData.plot_0.99.12 SpatialData.data_0.99.10 spatialdataR_0.99.47
[4] patchwork_1.3.2 ggplot2_4.0.4 dplyr_1.2.1
[7] sf_1.1-3
loaded via a namespace (and not attached):
[1] tidyselect_1.2.1 grumpy_0.2.0
[3] blob_1.3.0 farver_2.1.2
[5] filelock_1.0.3 R.utils_2.13.0
[7] S7_0.2.2 fastmap_1.2.0
[9] SingleCellExperiment_1.35.2 BiocFileCache_3.3.0
[11] duckdb_2.0.0 digest_0.6.39
[13] lifecycle_1.0.5 paws.storage_0.11.0
[15] RSQLite_3.53.3 magrittr_2.0.5
[17] compiler_4.6.2 rlang_1.3.0
[19] tools_4.6.2 yaml_2.3.12
[21] knitr_1.52 S4Arrays_1.13.2
[23] htmlwidgets_1.6.4 bit_4.6.0
[25] classInt_0.4-11 curl_8.1.0
[27] reticulate_1.47.0 DelayedArray_0.39.8
[29] xml2_1.6.0 RColorBrewer_1.1-3
[31] abind_1.4-8 KernSmooth_2.23-27
[33] withr_3.0.3 purrr_1.2.2
[35] BiocGenerics_0.59.12 R.oo_1.27.1
[37] grid_4.6.2 stats4_4.6.2
[39] e1071_1.7-17 scales_1.4.0
[41] SummarizedExperiment_1.43.0 cli_3.6.6
[43] rmarkdown_2.32 generics_0.1.4
[45] cachem_1.1.0 DBI_1.3.0
[47] proxy_0.4-29 BiocManager_1.30.27
[49] XVector_0.53.0 matrixStats_1.5.0
[51] vctrs_0.7.3 Matrix_1.7-6
[53] jsonlite_2.0.0 IRanges_2.47.5
[55] S4Vectors_0.51.10 bit64_4.8.6
[57] RBGL_1.89.0 ggnewscale_0.5.2
[59] units_1.0-1 glue_1.8.1
[61] ZarrArray_1.1.7 gtable_0.3.6
[63] Rarr_2.1.69 GenomicRanges_1.65.4
[65] tibble_3.3.1 pillar_1.11.1
[67] htmltools_0.5.9 Seqinfo_1.3.2
[69] graph_1.91.0 dbplyr_2.6.0
[71] R6_2.6.1 httr2_1.3.0
[73] wk_0.9.5 evaluate_1.0.5
[75] lattice_0.23-1 Biobase_2.73.2
[77] R.methodsS3_1.8.2 png_0.1-9
[79] duckspatial_1.4.0 memoise_2.0.1
[81] renv_1.3.1 paws.common_0.9.0
[83] class_7.3-24 uuid_1.2-2
[85] Rcpp_1.1.2 SparseArray_1.13.4
[87] anndataR_1.3.5 xfun_0.62
[89] MatrixGenerics_1.25.0 pkgconfig_2.0.3 References
Schapiro, Denis, Artem Sokolov, Clarence Yapp, et al. 2022. “MCMICRO: a scalable, modular image-processing pipeline for multiplexed tissue imaging.” Nature Methods 19 (3): 311–15. https://doi.org/10.1038/s41592-021-01308-y.